Open Vector Editor
Specs
| Label | Value |
|---|---|
| Version | 3.0.0 (updated on 2025-05-28) |
| Developer | Labii Inc. |
| Type | Section |
| Support Configuration | No |
Overview
The Open Vector Editor widget integrates TeselaGen's Open Vector Editor (OVE) directly into Labii so users can view, design, edit, and annotate plasmids and DNA sequences inside ELN and LIMS records. It supports circular and linear sequence views, feature annotations, cut-site inspection, virtual digests, and alignment workflows in one interactive editing environment. This widget is well suited for molecular cloning, plasmid verification, synthetic biology, and sequence documentation tasks where scientists need both a graphical map and sequence-level editing tools. By keeping sequence design and analysis in the same record as the rest of the experiment data, the widget improves traceability and collaboration.
Use Cases
- Plasmid Design: Build new plasmid constructs by entering DNA sequences and annotating promoters, coding regions, tags, and markers.
- Sequence Verification: Review imported plasmid files, inspect cut sites, and validate annotated features before downstream experiments.
- Sanger Verification: Align AB1 sequencing reads to the expected construct and inspect the chromatogram at each mismatch.
- Primer Design: Design PCR primers for cloning, colony PCR, or sequencing directly on the construct, and identify unknown regions with BLAST.
- Cloning Workflows: Simulate restriction digests and manage sequence edits during cloning and assembly planning.
- Inventory Documentation: Store plasmid sequences and annotations alongside inventory, storage, and usage records in Labii.
- Annotation Automation: Apply feature libraries or primer libraries to sequences to accelerate repetitive annotation tasks.
Interface
Read-only View
In read-only mode, the widget presents the saved sequence record with a reduced toolbar that contains review tools only. Users can review the plasmid or sequence map, sequence annotations, and other saved analysis outputs as part of the record. This view is useful for record review and handoff because it keeps the documented sequence visible while preventing accidental modification. Read-only mode applies when the record is not editable by the current user and when the record is opened from a share link.
- Data Display: Displays the saved sequence map and associated annotations.
- Annotation Layers: Show or hide cut sites, features, primers, and open reading frames, or use Show Visibility Options to choose which annotation layers are displayed.
- Export Options: Use Export to download the sequence, or Print Vector to print the map.
- Search/Filter: Use the find tool to search the sequence and annotations, and filter cut sites by enzyme or enzyme group.
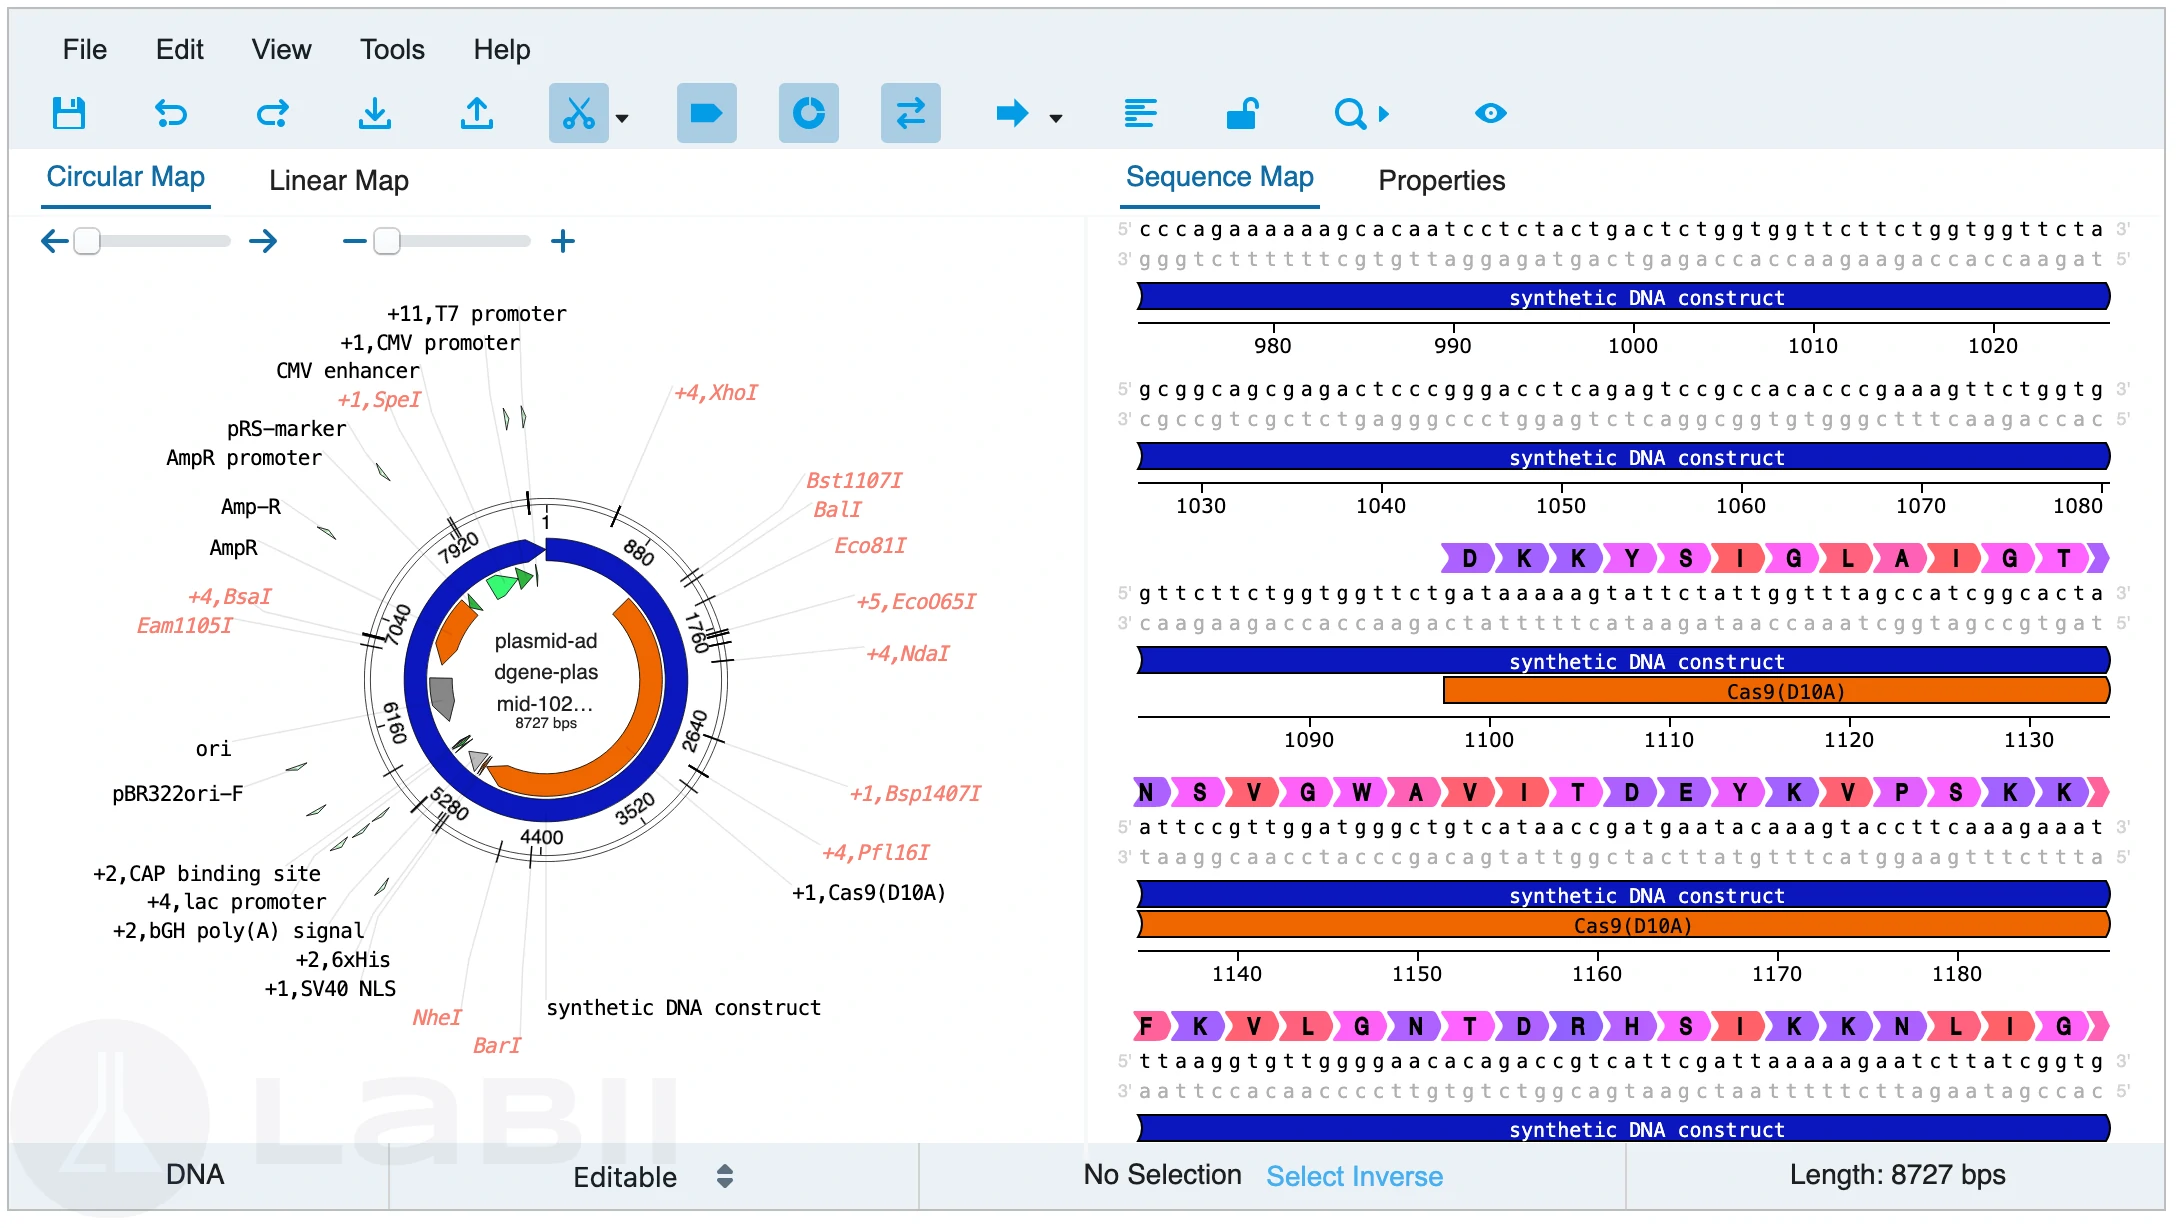
The read-only view displays the saved sequence record and annotations without the editing controls.
Edit View
In edit mode, the full OVE workspace opens so users can import sequences, switch between circular and linear views, annotate sequence features, inspect cut sites, and run supporting analysis tools. The main workspace combines a graphical map, a sequence viewer, and supporting panels for properties, features, and analysis outputs. This layout lets users move between visual construct editing and base-level sequence operations without leaving the record.
- Input Methods: Paste raw sequence text or import supported sequence files such as GenBank, ApE, FASTA, AB1, SnapGene, and related formats.
- Formatting Options: Switch among circular, linear, and other supported sequence views while displaying annotations and digest information.
- Validation: Inspect annotations, alignment outputs, and cut sites before saving changes back to the record.
- Status Bar: Shows the current selection and its percent GC content. Turn the GC display on or off in View > Percent GC Content of Selection.
- Collaboration: Save updated sequence records into Labii so other team members can review the latest construct state.
Configuration
The Open Vector Editor widget does not require widget-level configuration before use. Once the widget is added to a section, users can immediately load or enter sequence data and begin editing.
Because the widget has no separate settings panel, behavior is driven by the sequence data loaded into the editor and the analysis tools the user chooses during editing.
Additional Functions
Import Sequence
Open Vector Editor supports importing sequence data from common molecular biology file formats, including .gb, .gbk, .ape, .ab1, .fasta, .fas, .json, .xml, .seq, .sbd, and .dna.
Open the record and click Edit on the Open Vector Editor section.
Click Open in the toolbar, or drag a supported file onto the cloud icon.
Select the sequence file to import.
Review the loaded construct in the editor and save the record when finished.
Export Sequence
Sequences can be exported for sharing or downstream analysis in formats such as GenBank, FASTA, and TeselaGen JSON. Export is available in both edit and read-only modes.
Open the record that contains the Open Vector Editor section.
Click Export in the toolbar, or open File and select Export Sequence.
Choose the desired export format and download the file.
Print the Map
Print the current sequence map for lab notebooks, bench references, or review meetings. Printing is available in both edit and read-only modes.
Click Print Vector in the toolbar, or open File and select Print.
Confirm the print settings and print or save the map as a PDF.
Create a Sequence From a Selection
Save a region of the current construct, such as an insert or an expression cassette, as a separate GenBank file. Features inside the selected region are kept in the new file.
Select the region in the sequence map or sequence view.
Open Edit > Create and select Create New DNA Sequence From Selection. You can also right-click the selection and open Create.
The region downloads as a GenBank (.gb) file named after the section and the selected positions, for example pUC19_101-850.gb.
The new sequence is downloaded as a file. It is not added to the current record. To keep it in Labii, import the file into another Open Vector Editor section.
BLAST a Selection
Search a region of the construct against NCBI databases to identify an unknown insert, confirm a coding sequence, or check where a sequence comes from. This action is available in both edit and read-only modes.
Select the region in the sequence map or sequence view.
Right-click the selection and select BLAST Selection at NCBI, or open Tools and select BLAST Selection at NCBI.
The NCBI nucleotide BLAST page opens in a new browser tab with the selected sequence filled in. Choose the database and other search options on that page, then click BLAST.
This action sends the selected sequence to NCBI, an external public service. Do not BLAST confidential sequences unless your organization allows it. Selections longer than 7,000 bp cannot be sent by link; copy the sequence and paste it on the NCBI BLAST page instead.
Design Primers
Design a PCR primer pair for a selected region and add it to the sequence as primer annotations. Melting temperatures use the same SantaLucia method as Primer3.
Open the widget in edit mode and select the target region.
Right-click the selection and select Design Primers for Selection, or open Tools and select Design Primers for Selection.
Choose the Primer Placement:
- At the selection ends (amplify the selection): Primers start exactly at the ends of the selection, which is useful for amplifying an insert for cloning. Only the primer length is adjusted.
- Outside the selection (screen or sequence across it): Primers are placed outside the selection, within the search window, which is useful for colony PCR or for sequencing across an insert.
Review Minimum Length (nt), Maximum Length (nt), Target Tm (°C), and Search Window Outside Selection (bp). The defaults are 18–25 nt, 60 °C, and 200 bp.
Click Submit. The best primer pair is added to the sequence, named after the section with _F and _R, and a message shows the melting temperature of each primer and the product size.
Review any warning message, check the primers on the map, and run Tools > Simulate PCR to confirm the product. Click Undo to remove the pair if it is not suitable.
Primer pairs are scored on how close each melting temperature is to the target, the difference between the forward and reverse melting temperatures, GC content between 40% and 60%, a 3′ GC clamp, runs of five or more identical bases, 3′ self-complementarity, possible primer dimers, and whether the 3′ end binds more than one site in the sequence.
Primer design checks binding only within the current sequence. It does not check for off-target binding elsewhere in a genome, and only the best-scoring pair is added. Confirm critical primers with a dedicated tool before ordering.
Change Sequence and Features
The widget supports direct sequence editing together with feature creation, editing, and deletion.
Load or paste a sequence into the editor.
Select a sequence region in the editor.
Use the toolbar to insert bases, edit selected bases, delete selected bases, copy sequence, or create and update a feature.
Review the updated map and sequence view, then save the record.
Restriction Digest and Cut Sites
OVE includes enzyme and cut-site tools for planning cloning workflows and evaluating sequence manipulation strategies. By default, the map shows single cutters, which are enzymes that cut the construct exactly once.
Open the sequence in edit mode.
Click the arrow next to the cut site tool (Show Cut Site Options), or open Tools and select Filter Cut Sites, then choose the enzymes or enzyme groups to display.
Open Tools and select Simulate Digestion to view the fragments and a virtual gel for the selected enzymes, or Simulate PCR to preview a PCR product.
Review recognition sites, cut positions, and fragment information for the current construct.
Common Enzyme Groups
Labii adds built-in enzyme groups to the cut site filter, so you can show a standard set of enzymes without selecting each one.
| Group | Enzymes | Typical use |
|---|---|---|
| Common cloning enzymes | 64 | 6-bp cutters found in common multiple cloning sites, such as EcoRI, BamHI, HindIII, NotI, XhoI, NdeI, and NcoI |
| Golden Gate (Type IIS) | 11 | Enzymes that cut outside their recognition site, such as BsaI, BsmBI, Esp3I, BbsI, SapI, and PaqCI |
| Rare cutters (8+ bp) | 11 | Enzymes with long recognition sites, such as NotI, AscI, PacI, PmeI, SfiI, and SwaI |
| Frequent 4-cutters | 16 | Diagnostic and methylation-sensitive enzymes, such as DpnI, MspI, HpaII, Sau3AI, AluI, and HaeIII |
Open the cut site filter from Show Cut Site Options or Tools > Filter Cut Sites.
Select one or more enzyme groups. Groups can be combined with individual enzymes and with Single cutters.
Review the cut sites on the map, then run Simulate Digestion if you need fragment sizes.
Groups you create in Tools > Manage Enzymes appear in the same list as the built-in groups. Your own groups are stored in your browser, so they are not shared with other users or devices.
Auto Annotate
Labii supports automated annotation in OVE using CSV feature libraries or ApE feature files so users can quickly label commonly used sequence elements.
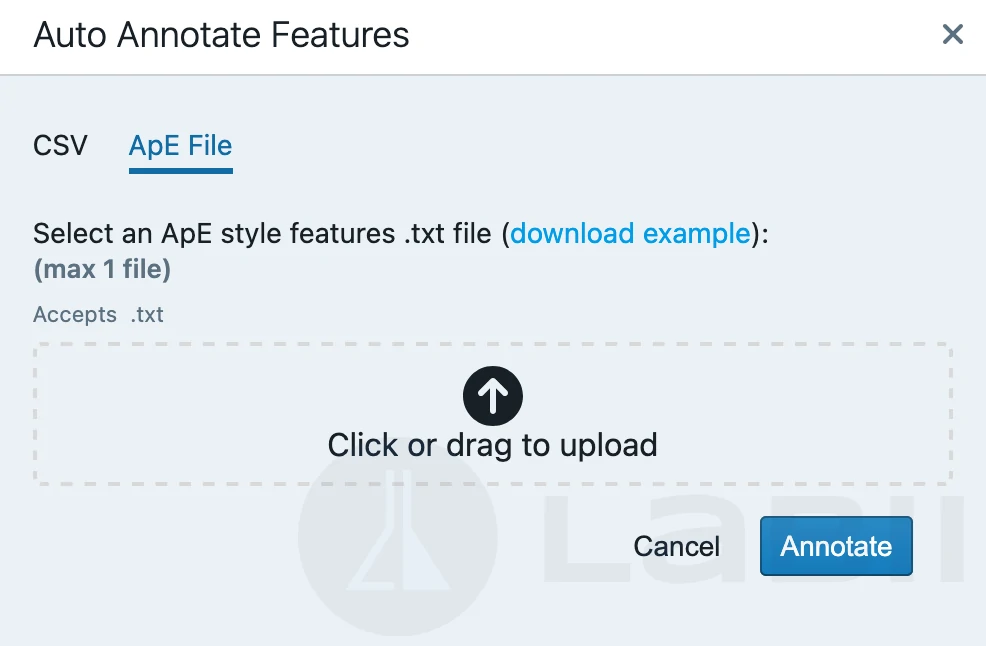
The auto-annotation workflow matches sequence regions against uploaded feature libraries and applies annotations automatically.
In the editor toolbar, open Tools and select Auto Annotate.
Choose whether to auto annotate features, parts, or primers.
Upload or select a CSV or ApE feature file.
Review the generated annotations and adjust them manually if needed.
Sequence Alignment
The alignment tool compares the active sequence against pasted sequences or uploaded sequencing reads so users can review sequence similarity and confirm construct identity. Upload Sanger sequencing results (.ab1) to verify a clone against its reference: the chromatogram trace and quality scores are shown under each read.
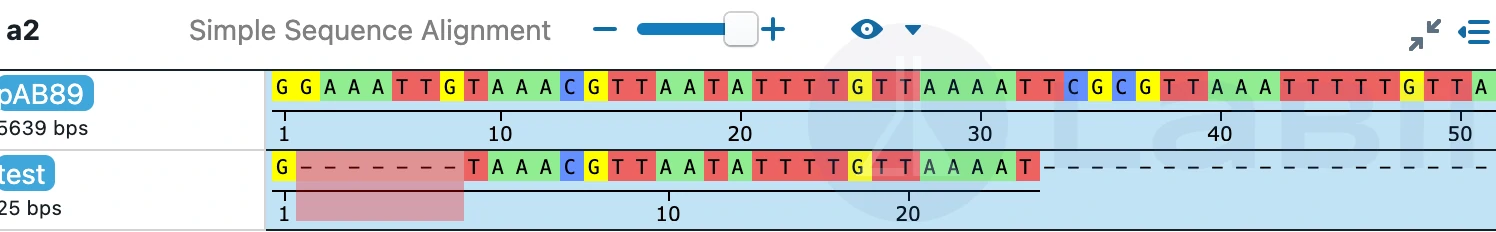
The alignment panel shows the comparison between the current construct and an input sequence for verification and review.
Open the widget in edit mode and click the Alignment icon in the toolbar.
Enter an Alignment Name.
Add the sequences to compare. You can use either option or both:
- Enter a Sequence Name and paste the sequence in Sequence. Select Should Reverse to align its reverse complement.
- Under Sequencing Reads (.ab1, .fasta, .gb), select one or more files, such as the forward and reverse reads for a clone.
Keep Trim low-quality ends selected to remove the low-quality bases at the start and end of each AB1 read before aligning.
Click Submit to generate the alignment. Uploaded reads are automatically reverse-complemented when they match the opposite strand, so all reads are shown in the same orientation as the reference.
Review the alignment panel, including the chromatogram peaks at any mismatch, and save the record to keep the result.
Alignment runs in your browser, and sequences are not sent to an external service. Each AB1 read saved with an alignment adds its trace data to the record, so align only the reads you need to keep.
Reads that span the origin of a circular plasmid are aligned as if the plasmid were linear and may show a large gap. Rotate the reference so the region of interest does not cross the origin before aligning.
Fullscreen and Standalone Review
Users can enlarge the editor interface to focus on sequence review and editing without distraction from other record sections.
Open the widget in edit mode.
Use Fullscreen or Standalone from the available interface actions.
Review or edit the construct in the expanded workspace, then save changes back to the record.
Best Practices
Sequence Management
- Import authoritative sequence files, such as curated GenBank or SnapGene exports, whenever possible instead of relying on manually retyped sequences.
- Use clear construct names and feature labels so downstream users can understand the plasmid record without reopening raw source files.
- Save sequence changes after each major editing step to keep the record synchronized with the current design.
Annotation Quality
- Review auto-generated annotations before accepting them as final, especially for overlapping features or ambiguous matches.
- Standardize feature naming across projects so searches, comparisons, and exports remain consistent.
- Confirm strand direction, start and end positions, and feature type before finalizing a plasmid map.
Workflow Reliability
- Check cut sites and virtual digest results before ordering primers, enzymes, or cloning reagents.
- Use the alignment tool to verify sequence identity when comparing expected and observed constructs.
- Prefer the same editor across related projects when consistency of visualization and annotation behavior matters.
Using imported reference files, standardized annotations, and saved intermediate edits helps maintain reliable plasmid records and reduces rework during cloning and verification.
Common Pitfalls to Avoid
- Avoid saving a construct without reviewing imported annotations, because imported files may contain outdated or inconsistent feature labels.
- Avoid making large sequence edits without rechecking cut sites and alignments afterward.
- Instead verify the edited construct with digest and alignment tools before treating the record as final.
Related Widgets
- Plasmid Editor: Use this widget when you need another plasmid-focused editing workflow with sequence map and feature management inside the same molecular biology suite.
References
- Open Vector Editor on GitHub - Source project for the embedded OVE editor.
- Molecular Biology Suite - Category overview for related sequence analysis widgets in Labii.
- ApE Feature Files - Reference format supported by the auto-annotation workflow.